Home  /  Quality

ISO 13485 quality management system.

Certification establishes that the system was audited. Supplier qualification requires more: this page sets out system scope, the records produced, and the audit access available prior to engagement.

Single-site operation. The process under audit is the process in daily use.

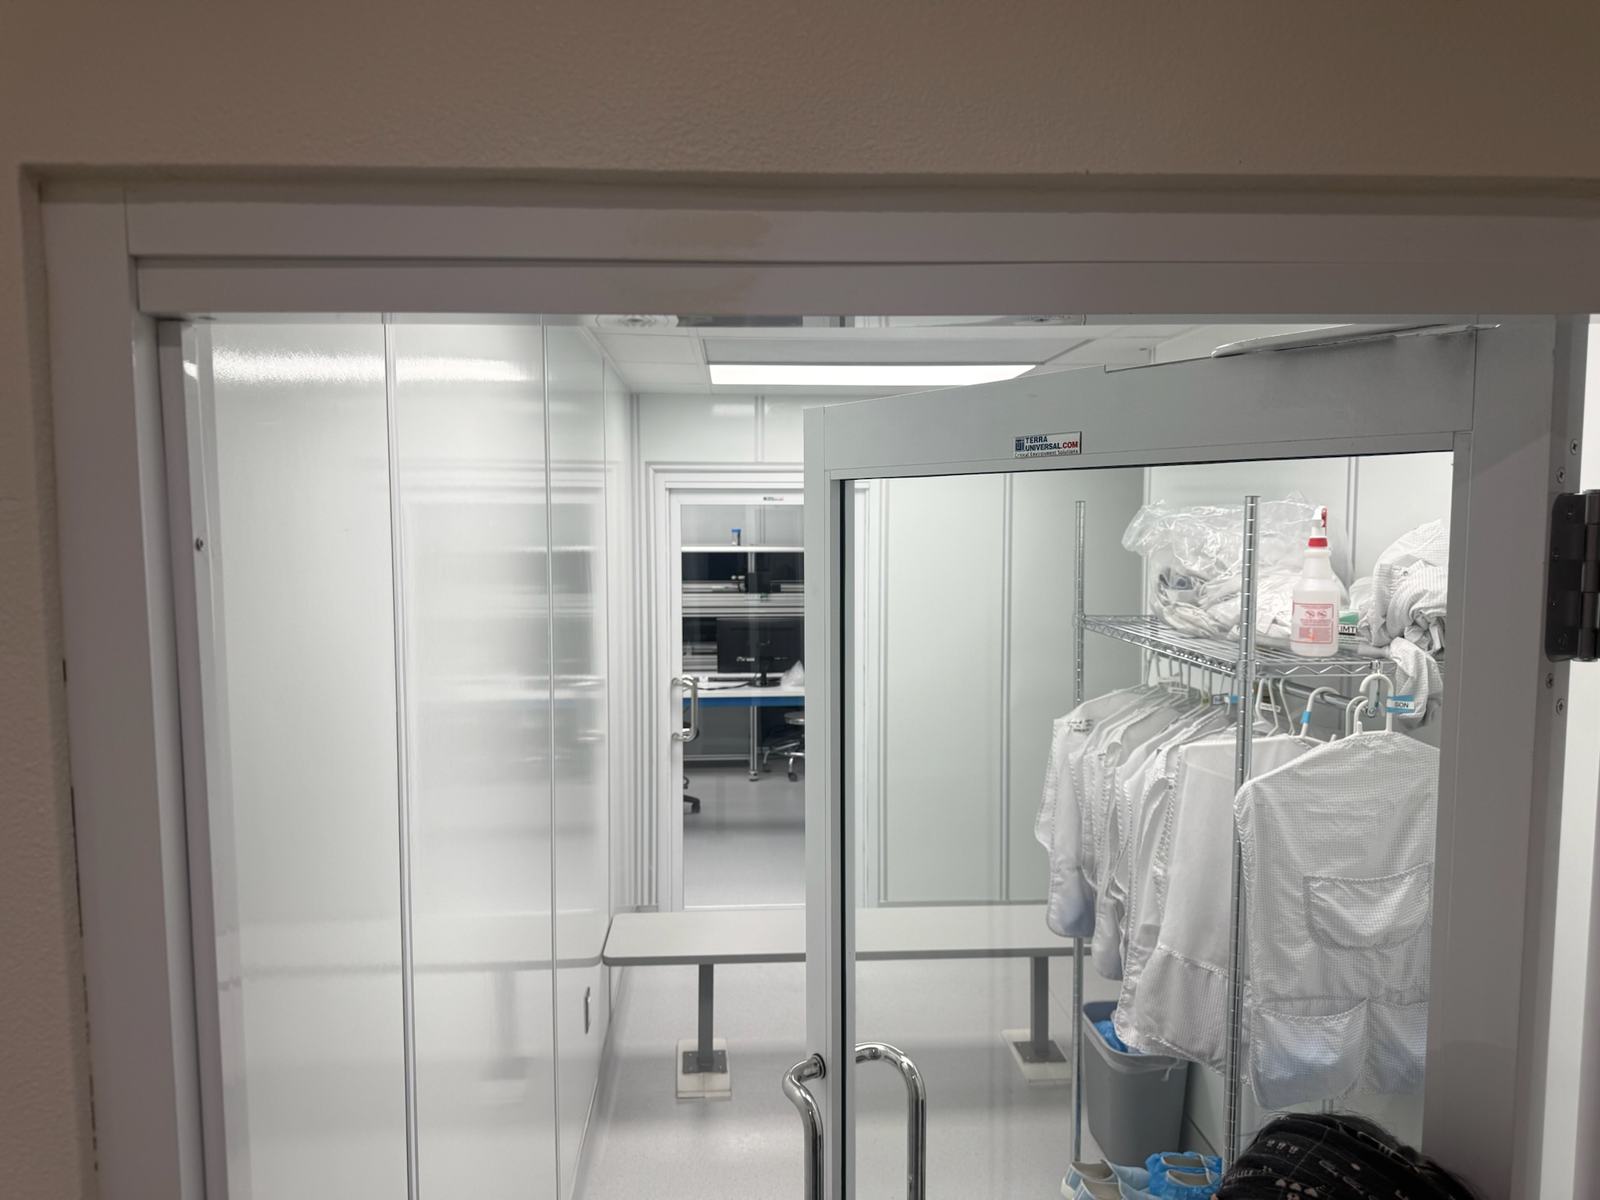
Gowning room with coveralls staged before entry to the MedTech Innovation cleanroom
Quality system
ISO 13485Medical devices
Cleanroom
Class 10,000ISO 7, gowned, positive pressure
Records
YoursDMR and design history belong to you
Audits
ScheduledAvailable prior to engagement
Certifications Quality policy The system Records Whose QMS Cleanroom control Audit us

Certifications & registrations

Certifications and registrations.

Registrar, certificate number and expiry date are stated below for independent verification. Certificates are available on request.

Held

ISO 13485 — Quality Management System

Medical device quality management system covering design, development and manufacture, to the standard your own quality organization audits against.

Held

Class 10,000 / ISO 7 cleanroom

Hard-wall modular construction with sealed epoxy flooring, gowning, a transfer airlock and filtered positive-pressure downflow. ISO 7 permits roughly ten times fewer airborne particles than ISO 8.

Only credentials currently held are listed.

Issued and posted on site

Quality policy.

Quality policy

MedTech Innovation, LLC will consistently meet and strive to exceed customer expectations through the manufacture and sale of high-quality products and by providing high-quality customer service. MTI is committed to complying with quality requirements, to maintaining the effectiveness of the quality management system, and to periodically reviewing the quality system for continuing suitability.

What the system covers

System elements and program touchpoints.

Required elements of a certified system. Listed below with the point at which each becomes visible during an active program.

ElementWhere it touches your program
Design controlsDesign inputs, outputs, reviews and traceability through the five-phase program, so a design freeze is a documented state rather than a feeling.
Risk managementRisk assessment and FMEAs created in design and updated as the design and process lock down through design for manufacturing.
Document controlDrawings, specifications and work instructions under revision control, so the technician at the bench is building to the version you approved.
Record retentionBuild records, inspection records and test results captured as the work happens and retained, not reconstructed afterward.
Supplier controlsComponent vendors qualified and managed during design for manufacturing, so the supply chain is finalized before the V&V build, not during it.
Incoming inspectionMaterials received and documented against acceptance criteria, so a discrepancy surfaces before it is built into product.
Training recordsEvery technician who touches your device is trained and recorded as trained — including gowning and material handling for anyone entering the cleanroom.
Corrective & preventive actionNonconformances investigated and closed, with the record available to you rather than filed somewhere you never see.
Change controlChanges after a design freeze go through a documented path. This is the mechanism that stops "one small tweak" from quietly invalidating a V&V build.
Management reviewThe quality system reviewed periodically for continuing suitability, per the policy above.

The paper trail

Documentation chain.

Each record is created as a precondition of the next. This chain constitutes the file reviewed by a regulator, auditor or acquirer.

Documentation chainEverything here belongs to you
01

Design inputs

What the device has to do, written down before anyone builds to it.

02

Design outputs

Drawings, specifications and the risk file that answer those inputs.

03

Device master record

The complete recipe for the device — what it is made of and how it is made.

04

Work instructions

The DMR translated into steps a trained technician executes the same way every time.

05

Device history record

Proof of what was actually done to each build: who, when, with which lot.

06

Release

Inspected, tested, packaged, seal-verified and released against the record.

Records and materials are returned at engagement close-out. The device master record and design history remain customer property irrespective of staffing model.

The question rental raises

Applicable quality system by staffing model.

Two defined arrangements, determined by which organization staffs the benches.

Your team staffs the benches

Your quality system

You run your process, under your procedures, producing your records. We provide the controlled environment and the facility controls that support it — and evidence of those controls for your file.

  • You own the process, the records and the release decision
  • We supply environmental and facility control evidence
  • Your people are trained to our facility procedures for entry, gowning and material handling
  • Your auditors audit you; we support with facility documentation
Our technicians staff the benches

Our ISO 13485 system

The build runs under our quality management system, to your approved documentation. Records are produced as the work happens and delivered to you.

  • Work performed to your approved drawings and specifications
  • Trained, recorded technicians under document and change control
  • Build and inspection records delivered with the product
  • You can audit the system that produced them, before or during

Mixed arrangements are common — our technicians early, yours later as you hire. When that happens the handover point is defined in writing before it occurs, not discovered in a record review afterward.

Environmental control

Contamination control is a sequence, not a room.

Reception, gowning, transfer airlock, Class 10,000 floor, packaged and sealed. Each step removes what the step before it could not, with filtered positive-pressure downflow so air always moves from cleaner space toward less clean space. Anyone entering is trained on the entry procedure and recorded as trained.

See the full cascade and facility →
GowningTrained and recorded before entry.
Transfer airlockMaterials wiped down and passed through.
Pressure cascadeFiltered positive-pressure downflow.

Before you commit

Facility and quality system audits.

Customer audits are scheduled in advance and conducted under NDA. Scope includes floor walkthrough, quality record review, and access to the technicians assigned to the work.

Single-site operation. The process presented at audit is the process in daily production use.

Available on request

Send a request and we will send what applies to your review. Under NDA where it needs to be.

  • ISO 13485 certificate
  • Cleanroom certification report
  • Signed quality policy
  • Capability statement
  • Facility and environmental control summary
  • Supplier questionnaire responses
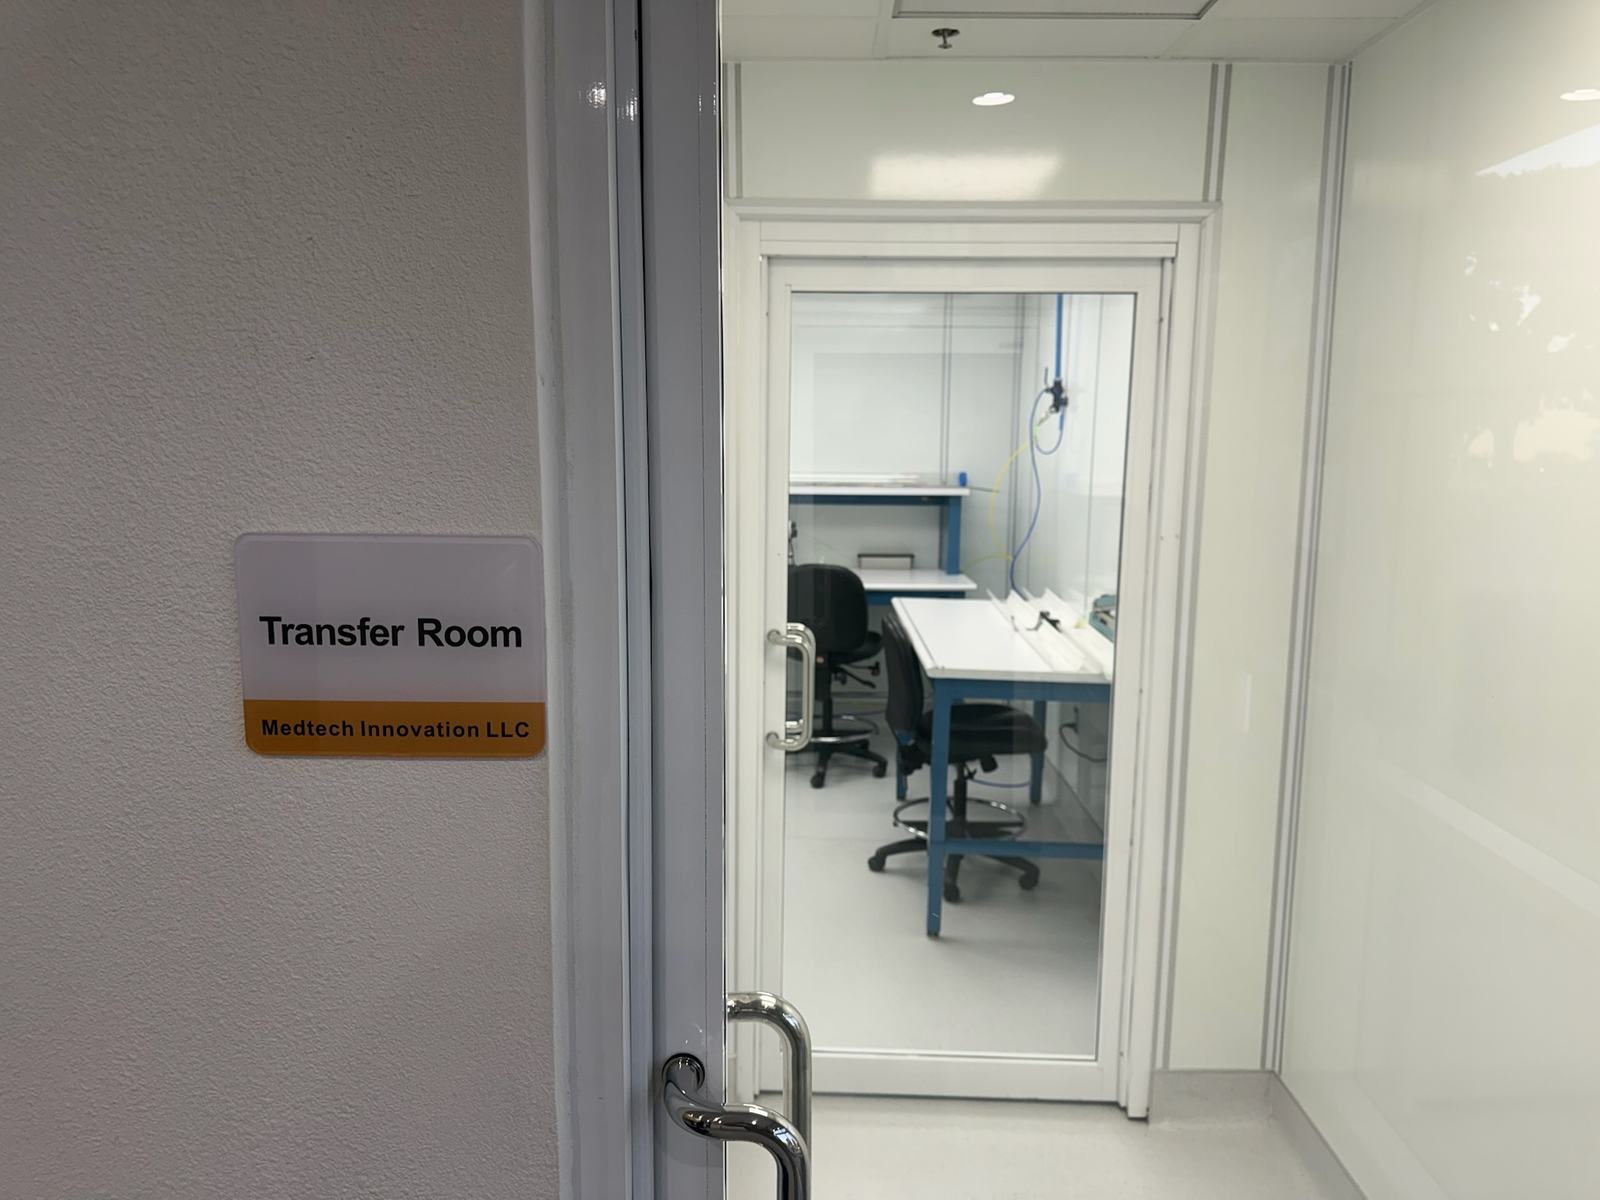
Transfer room airlock at the MedTech Innovation cleanroom
Transfer room — material control into the classified space

Honest boundaries

Services performed externally.

The following are performed by qualified partners or by the customer, and are identified at quotation.

Handled elsewhere, or by you

  • Sterilization — coordinated with third-party providers based on customer requirements
  • Biocompatibility testing — external lab
  • Specialized inspection and metrology
  • Tensile testing and flow measurement
  • Regulatory submissions — we support, we do not file for you
  • Clinical trial management

We support each of these — documentation, samples, coordination and build — but the work itself happens with a qualified partner or with your team.

Quality documents

Request quality documentation.

Certificates, quality policy, facility control summary and supplier questionnaire responses, as required for supplier qualification. On-site audits scheduled on request.

+1 949-336-2972 [email protected]

17151 Gillette Ave, Irvine, CA — 8 minutes from John Wayne Airport
Mon–Fri, 8:00 AM – 4:30 PM PT

Document requests are answered by the next business day.